GROMACS
engines/gromacs/seams_ice.cpp reads a trajectory in
analyzeFrame and multiplies coordinates by 10, from nanometers to
angstrom. engines/gromacs/ice.gro stores the second oxygen at
0.1588 nm.
gromacs atoms 64 scaled_A 1.5880 ice 64
The walkthrough checks that 0.1588 nm becomes 1.5880 angstrom before
CHILL+, and that the converted cell is 64 cubic molecules. When
gmx-seams is built it also has to print gromacs frames 2 ice 128
atoms 128, because the tool reads the gro twice.
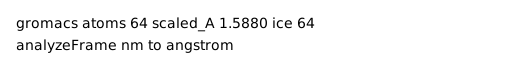
analyzeFrame converts the gro from nanometers.
What this tells you
The trajectory hook’s unit conversion is what makes 0.1588 nm an oxygen-oxygen contact. With that factor the frame is 64 cubic molecules.